Quick answer: Plan DQ, IQ, OQ and PQ as one traceable evidence chain for the configured capsule blister line. IQ confirms that the installed equipment matches the approved design and installation requirements. OQ challenges the machine and its controls across defined operating ranges. PQ demonstrates that the complete process can repeatedly produce acceptable capsule blisters under the buyer’s approved production conditions.
The validation boundary should cover the actual product, forming and lidding materials, tooling, capsule feeding, sealing, registration, cutting, inspection, rejection, data functions and downstream interfaces included in the project. The buyer remains responsible for the site quality system, process validation strategy and product release.

What do DQ, IQ, OQ and PQ mean for capsule blister packaging?
The four stages answer different questions. Treating them as interchangeable creates duplicated tests, missing evidence and arguments late in the project.
Design Qualification (DQ) connects the User Requirements Specification (URS) to the proposed machine, options, materials, tooling, inspection concept, software boundary and documents. Installation Qualification (IQ) records what was actually installed. Operational Qualification (OQ) challenges functions and operating ranges. Performance Qualification (PQ) uses the approved process, product and pack to show repeatable performance under the buyer’s protocol.
Keep capsule filling and blister packaging in separate validation scopes
A capsule filling machine and a capsule blister packaging machine perform different processes. The filler controls empty-capsule separation, dosing, closing and capsule-weight or integrity checks. The blister line receives finished capsules and controls their presentation, cavity formation, placement, sealing, inspection, coding and finished-card discharge.
If both systems are part of one project, define the handoff explicitly: capsule condition, orientation, dust level, allowable damage, bulk container, transfer method, accumulation, inspection status and data ownership. Qualification evidence from the filler does not automatically prove performance of the blister line, and blister-line evidence does not replace the filling process validation.
Scope rule: qualify the interface as well as each machine. A stable upstream average can still hide broken capsules, joined capsules, excessive dust or inconsistent presentation that disrupts downstream feeding.
Build the validation lifecycle before the purchase order
The best time to define evidence is before supplier selection. A quotation that says “validation documents included” is not enough. Name the documents, language, format, revision workflow, approval responsibility, execution location, test materials, acceptance criteria, raw-data ownership and support boundary.
| Stage | Primary input | Evidence to retain | Decision enabled |
|---|---|---|---|
| URS and risk review | Product, pack, output, quality and data requirements | Approved URS, risk inputs, scope and exclusions | What must the project prove? |
| DQ | Supplier design response | URS traceability, drawings, specifications and open items | Is the design suitable to proceed? |
| FAT | Approved factory test protocol | Configuration, settings, results, rejects, stops and deviations | Is the equipment ready for shipment? |
| SAT / commissioning | Installed site conditions | Utilities, interfaces, safety checks and commissioning records | Is the system ready for qualification? |
| IQ | Approved design and installed baseline | Component, utility, drawing, software and calibration records | Was the correct system installed? |
| OQ | Critical functions and operating ranges | Challenge results, alarms, interlocks and deviations | Can the system operate as intended? |
| PQ | Approved product and production process | Run records, samples, test results and disposition | Is performance repeatable in routine conditions? |
FDA’s process validation guidance describes validation as a lifecycle activity supported by scientific knowledge and continued assurance. EU GMP Annex 15 likewise places qualification and validation within a planned, risk-based lifecycle. Neither source turns a generic supplier document bundle into an approved site validation program. The buyer should interpret applicable requirements with its own quality and regulatory teams.
Define the capsule blister machine validation boundary
Start at the first controlled input and end at the last accepted output. For one project this may begin with bulk capsules and end with cut blister cards. For another it may include an upstream conveyor, print system, vision inspection, reject confirmation, cartoner interface and electronic batch data.
| System area | Qualification questions | Typical retained evidence |
|---|---|---|
| Material unwind and web control | Are reel direction, tension, tracking and splice handling defined? | Material identification, setup, settings and observations |
| Forming | Does the approved web form repeatable cavities without unacceptable damage? | Tooling revision, settings, cavity measurements and defect samples |
| Capsule feeding | Can representative capsules enter cavities without damage, doubles or misses? | Product lots, feeder settings, counts, defects and interventions |
| Sealing | Is there a usable operating window for the approved material pair? | Temperature, pressure, dwell or speed settings and integrity results |
| Registration, coding and cutting | Does the finished card match the approved drawing and artwork controls? | Printed samples, position measurements, code results and cut limits |
| Inspection and rejection | Are defined defects detected and rejected, with reject confirmation where required? | Challenge set, detection results, rejected samples and disposition |
| Controls and data | Are access, recipes, alarms, records, backup and audit functions within scope? | Functional tests, role matrix, records, exports and backup evidence |
| Line interfaces | Can upstream and downstream equipment exchange product and status safely? | I/O checks, accumulation tests, stop logic and interface records |
Write a testable URS and responsibility matrix
A strong URS states what must be achieved and how acceptance will be judged. Replace “high efficiency,” “GMP design” and “easy changeover” with measurable requirements tied to the intended product, materials and process. Each requirement should have an owner, verification method and retained record.
- Identify capsule sizes, shell materials, colors, dimensions, fragility, dust tendency and representative lots.
- Freeze the approved blister drawing, cavity orientation, forming web, lidding material, print registration and cut tolerances.
- Define accepted output at the discharge—not only mechanical cycles per minute.
- Name critical defects, inspection methods, reject logic and reject disposition.
- Define recipes, access levels, alarms, records, backup and electronic-data expectations in the contracted scope.
- State the planned FAT, SAT, IQ, OQ and PQ responsibilities and which party approves each protocol and report.
A responsibility matrix should cover the buyer, machine supplier and any third party. It should identify who supplies product and packaging materials, who authors and approves protocols, who calibrates instruments, who executes tests, who owns deviations, who archives raw data and who performs site re-testing after installation.
IQ checklist for a capsule blister packaging machine
IQ establishes the installed baseline. It should verify the agreed configuration, not simply repeat a packing list.
- Equipment, modules, serial numbers and nameplates
- Approved layout, drawings and installation location
- Product-contact and specified construction materials
- Forming, sealing and cutting tooling identification
- Electrical, pneumatic, cooling and extraction utilities
- Safety guards, interlocks and emergency stops
- Installed software or firmware versions in scope
- Measuring devices and calibration status
- Manuals, drawings, certificates and spare-parts lists in scope
- Cleaning, lubrication and preventive-maintenance information
- Backup media, recipe baseline and access-role list
- Open items, deviations and approved disposition
Record the “as installed” state at the time of execution. If a component, software version or utility connection changes later, assess the effect through change control instead of silently editing the old IQ record.
OQ should challenge functions, ranges and failure responses
OQ is more than a demonstration at one nominal setting. Use risk assessment and development knowledge to select meaningful challenges around the approved operating region. The protocol should define prerequisites, instruments, sampling, expected results, actual results and deviation handling before execution.
| OQ decision | Example challenge | Evidence to retain | Question answered |
|---|---|---|---|
| Web handling | Approved reel, tracking and controlled splice condition | Settings, observations, alarms and result | Can the exact web feed consistently? |
| Forming repeatability | Defined operating range and representative sampling | Cavity measurements, defect log and approved samples | Does the web form the approved geometry? |
| Capsule feeding | Representative capsule variants and controlled interruptions | Misses, doubles, damage, stops and interventions | Can the feeder handle the product range? |
| Seal process | Approved material pair across justified conditions | Recorded settings and agreed integrity-test results | Is there a usable sealing window? |
| Registration and cutting | Print-mark, code, perforation and cut-position checks | Measured samples and acceptance limits | Does the finished card match its controls? |
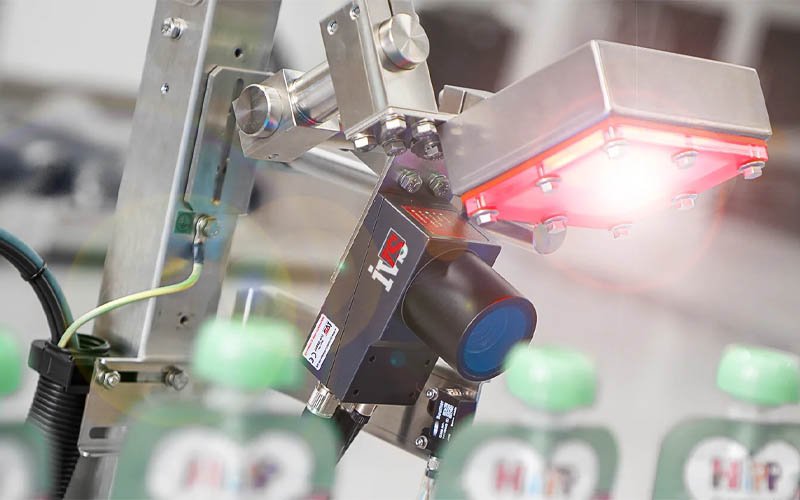
| Inspection and rejection | Approved good and defective challenge set | Detection, rejection, confirmation and disposition results | Are defined defects segregated? |
| Alarms and recovery | Selected sensor, guard, utility and stoppage conditions | Alarm records, system response and recovery steps | Does the line fail and recover as intended? |
Do not invent universal OQ limits
Temperature, pressure, dwell, speed and inspection thresholds depend on the product, materials, pack drawing, tooling and test method. The supplier can provide equipment knowledge and trial data, but the approved protocol should state project-specific conditions and acceptance criteria. A catalogue range is not automatically a validated operating range.
PQ proves the complete process under approved production conditions
PQ should use the approved machine configuration, trained operators, released materials, representative capsules, defined sampling and approved test methods. Its purpose is to show that the process can repeatedly produce packs that meet predefined requirements. The protocol should explain how batches, lots, shifts, operators, formats and worst-case conditions are selected.
There is no single run count that fits every capsule blister project. The buyer’s quality unit should justify the study design from process knowledge, risk, variability and applicable requirements. The machine supplier should not promise that a fixed number of runs automatically establishes validation.
| PQ input | Define before execution | Retained result |
|---|---|---|
| Product and lots | Capsule size, shell, fill characteristics, appearance and representative lot selection | Identity, status, quantities and lot traceability |
| Packaging materials | Forming and lidding grades, suppliers, widths, print and approved lots | Material identity and certificates or release status in scope |
| Operating condition | Recipe, target settings, line interfaces, duration and planned interventions | Settings, start/stop times, alarms and changes |
| Sampling and tests | Locations, frequency, quantity, instruments and acceptance criteria | Raw data, test results and sample disposition |
| Output accounting | Total packs, accepted packs, rejects, waste and rework rules | Reconciled counts and accepted-output result |
| Deviation handling | Recording, investigation, impact assessment and approval path | Deviation record, action and final disposition |
Challenge the process where capsule blisters actually fail
Capsule feeding and handling
Capsules can vary in dimensions, static behavior, surface condition and mechanical strength. Confirm the acceptable product condition at the feeder inlet, then record misses, doubles, damaged capsules, bridged product, stoppages and manual interventions. Use actual or justified representative products rather than assuming all capsule sizes behave alike.
Forming and cavity protection
The cavity must protect the capsule without creating excessive clearance or contact. Verify cavity geometry, wall condition, forming repeatability and web tracking using the approved drawing and material. Thermoform and cold-form routes require different forming knowledge; do not transfer settings or acceptance limits between them without evidence.
Sealing and package integrity
Control the material pair, seal area, product clearance, temperature, pressure, dwell or line speed and cooling conditions relevant to the design. Use the buyer-approved integrity or seal test and sampling plan. A visually acceptable seal alone may not answer the project’s protection requirement.
Inspection, rejection and output
Define defect classes before testing. Depending on scope, challenges may include missing, doubled, damaged or misplaced capsules; forming defects; lidding or registration issues; code failures; and reject-system faults. Verify not only detection but also physical removal, confirmation and controlled disposition. Accepted output should exclude rejected packs, planned samples, waste and unapproved rework.

Include cleaning, changeover and electronic functions where they matter
Cleaning and changeover are not optional side topics when they affect contamination control, mix-up prevention or repeatability. Define parts to remove, hard-to-reach areas, cleaning tools, line clearance, format-part identification, reassembly checks and the status of residual product. The buyer determines whether equipment cleaning supports a separate cleaning-validation program.
For electronic functions, define the intended use before testing. Recipe selection, role-based access, alarms, audit trails, electronic records, data export, backup and restore may require additional assessment when they are used for regulated decisions. A standard HMI does not by itself establish compliance. The buyer should map applicable data-integrity and computerised-system requirements to the actual configured functions.
Sampling, deviations and statistics need an approved method
A protocol should state where samples are taken, when they are taken, how many are tested, what instruments are used and how results are evaluated. Sampling at the beginning, middle and end of a run may be useful, but it is not automatically sufficient. Align the plan with identified variability, failure modes, process duration and the decisions the study must support.
Record deviations when they occur. Do not delete a failed result because a later adjustment passed. The final report should distinguish planned changes, unplanned interventions, invalid tests, confirmed failures, corrective actions and retesting. It should explain the impact on the conclusion and retain the approval trail.
Specify the document package in the quotation and contract
Name each deliverable rather than buying a vague “validation package.” The exact list depends on the project, but the commercial scope should address:
- URS response and scope/exclusion list
- Approved configuration and component list
- Drawings, manuals and certificates in scope
- Risk-review inputs and traceability matrix
- FAT protocol, raw results and final report
- IQ and OQ protocol templates or executed records
- Software, recipe and backup information in scope
- Calibration evidence for test instruments
- Deviation and open-item register
- Training and site-service records
- Change notification and support boundary
- Language, format, revision and approval workflow
The equipment supplier can provide machine documentation, configuration knowledge, trials and technical support. Those deliverables support the buyer’s qualification work; they do not replace the buyer’s pharmaceutical quality system, process validation, regulatory assessment or product-release responsibility.
Maintain the qualified state after approval
Qualification does not end with the final signature. Establish controls for recipes, tooling, software, sensors, inspection settings, packaging materials, maintenance, calibration and replacement parts. Assess changes before implementation and determine whether document updates, targeted requalification or broader validation work is needed.
Routine review should consider deviations, reject trends, recurring stops, maintenance history, calibration results, complaints and process changes. The aim is continued assurance that the approved process remains capable—not a one-time folder of signed documents.
Official references and related technical guides
- FDA: Process Validation—General Principles and Practices
- EU GMP Annex 15: Qualification and Validation
- EU GMP Annex 11: Computerised Systems
- Capsule blister packing machine configuration
- How a capsule blister packing machine works
- Common capsule blister packing problems
- 21 CFR Part 11 blister machine checklist
- Blister packaging materials guide
- Calculate real blister machine output
Frequently asked questions
What is IQ OQ PQ for a capsule blister packaging machine?
IQ verifies the installed machine and documentation against the approved design. OQ challenges functions, controls and operating conditions. PQ demonstrates repeatable finished-pack performance using the approved product, materials, process and sampling plan.
Is DQ required before IQ OQ and PQ?
DQ is commonly used to document that the proposed design addresses the URS and identified risks before installation. The buyer should define the applicable lifecycle and approval requirements in its validation plan and quality system.
Can the machine supplier execute IQ OQ and PQ?
The supplier may author templates, provide evidence, support execution or perform contracted tests. The buyer should define approvals, site responsibilities, deviation ownership and final acceptance. Supplier work does not transfer the buyer’s quality-system responsibility.
How many PQ batches are required for capsule blister packaging?
There is no universal number for every project. The buyer’s quality unit should justify the study design from process knowledge, risk, variability, applicable requirements and the intended commercial process.
What should be tested during OQ?
OQ should challenge the functions and conditions that can affect product handling, forming, sealing, registration, cutting, inspection, rejection, alarms, recovery and data functions within the approved scope.
Does FAT replace OQ?
No. FAT can generate valuable pre-shipment evidence and may reduce justified repetition, but site installation, utilities, interfaces and the buyer’s approved qualification strategy still need to be addressed.
Should vision inspection be included in qualification?
Include it when it is part of the configured control strategy. Define defect classes, challenge samples, detection criteria, rejection, reject confirmation, records and sample disposition before testing.
What information should I send for a validation-ready quotation?
Send capsule and pack details, material grades, blister drawing, accepted-output target, inspection and data requirements, line interfaces, document list, test expectations, site utilities and the proposed FAT, SAT, IQ, OQ and PQ responsibility boundary.
Define the evidence before the machine scope is frozen
Send the capsule samples, blister drawing, material grades, accepted-output target, inspection concept and validation boundary. HIJ can review the machine configuration, document list and FAT evidence needed for a testable project.
Discuss My Capsule Blister Validation Scope